🔥200名以上が申込済み
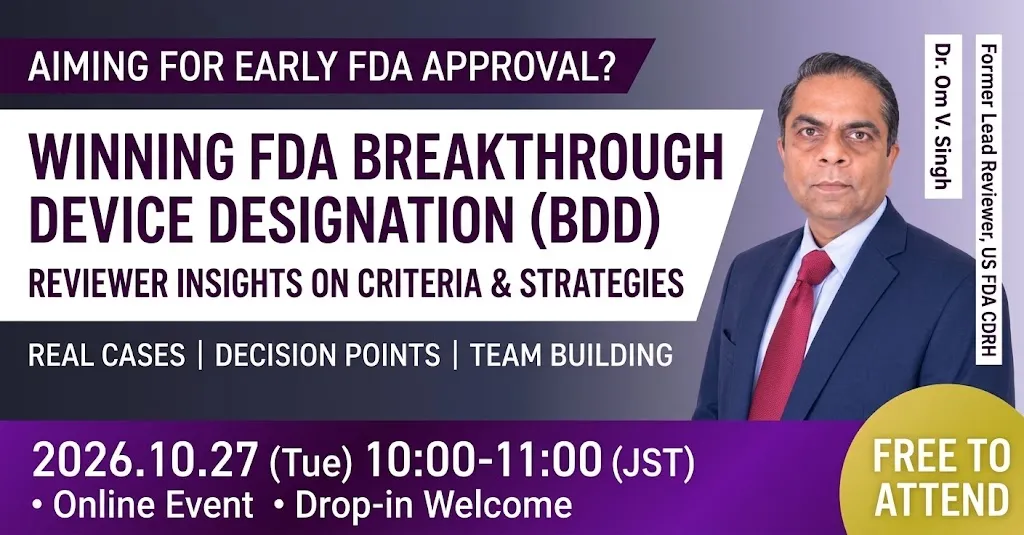
[Medical Devices / IVD] Seminar on submissions to the US FDA
Breakthrough Device Designation (BDD):
the evaluation criteria, and the team behind a submission — from a former FDA reviewer.
・Date & time: Tuesday, 27 October 2026, 10:00–11:00 (JST / UTC+9)
・Live on Zoom — your personal link is emailed to you right after you register
・Free to attend (a recording is sent to everyone who registers)
・Delivered entirely in English
・開催日時: 2026/10/27(火) 10:00〜11:00 (JST)
・Zoom開催
・無料セミナー(後日アーカイブ動画も配信)
・※本セミナーは英語で実施します。
Seminar overview
A Pre-Submission is an important opportunity to obtain the FDA's views on a focused set of questions, and further exchanges can follow as development progresses. But each one is a separate request.
Once a device receives Breakthrough Device Designation (BDD), the opportunity opens for more frequent, interactive communication with the FDA throughout development, and its subsequent submissions receive prioritized review.
· Sprint Discussions — time-bound, focused exchanges with the FDA on a specific issue
· Prioritized review of IDE, 510(k), De Novo and PMA submissions
· Involvement of FDA senior management and an experienced review team
· Flexibility to move part of the premarket data collection to the post-market phase
The bar, however, is not low. What distinguishes a strong BDD candidate is not innovation alone, but whether the device has the potential to provide more effective treatment or diagnosis for a life-threatening or irreversibly debilitating condition and can be supported by a clear, evidence-based rationale demonstrating that it meets the FDA's Breakthrough Device criteria.
And whether there is a team able to build that rationale.
Dr. Om V. Singh — a former Lead Reviewer at the US FDA's CDRH, still practising FDA consulting in the United States — will spend sixty minutes on the criteria as seen from the reviewing side, designated cases, how the rationale is built, and how to put the submission team together.
What attendees will take away
✔︎ What BDD is — the background to the program, and where it sits among the other consultation and submission routes
✔︎ What a designated sponsor actually receives — Sprint Discussions, prioritized review of IDE / 510(k) / De Novo / PMA, senior management involvement, flexible study design
✔︎ How to argue the eligibility criteria — effectiveness against a life-threatening or irreversibly debilitating condition, plus any one of the additional statutory criteria
✔︎ Three to five designated cases that can be traced in public information — company, therapeutic area, how the designation came about, and what followed
✔︎ When to consider it and who to work with — choosing between a Pre-Submission and a BDD, and building the team that constructs the rationale
Speaker
Dr. Om V. Singh
Former Lead Reviewer, US FDA CDRH / xCARE Expert
A regulatory and product development specialist with more than 25 years of experience across the US FDA, the medical device and pharmaceutical industries, and academia. He served as a Lead Reviewer at the US FDA's CDRH (Center for Devices and Radiological Health), and has held regulatory and scientific roles at Medtronic and UnitedHealth Group.
He specializes in regulatory strategy for medical devices, in-vitro diagnostics (IVD), SaMD and AI-enabled technologies, drug-device combination products and pharmaceuticals. His support covers BDD eligibility assessment and designation strategy; the 510(k), De Novo, PMA, IDE and Q-Submission routes; negotiation with the FDA; building clinical and non-clinical evidence; benefit-risk assessment; and lifecycle management from early development through to the post-market phase. He is also well versed in European regulatory strategy under the EU MDR and IVDR, and currently serves concurrently as a lecturer at Johns Hopkins University. Dr. Singh has published numerous research and review articles across multiple scientific disciplines and regulatory affairs.
Program
60 minutes in total — 45 minutes of presentation plus 10–15 minutes of Q&A
SESSION 1
What BDD is: the background to the program, and where it sits among the other routes
▶︎
What Breakthrough Device Designation is — when and against what background it began, and what the FDA intends it to do
▶︎
How it relates to the other consultation and submission options — Pre-Submission / Q-Sub / 510(k) / De Novo / PMA / IDE
▶︎
Breakthrough Device cases that are disclosed in public information — company, therapeutic area, how the designation or approval came about, and what followed (3–5 cases)
SESSION 2
The process up to a submission, and what begins once the designation is granted
▶︎
Not every company should apply — starting with where that line falls
▶︎
The process leading up to a submission, and how long it takes for a decision to come back
▶︎
How the relationship with the FDA may change after designation — the opportunity for more frequent, interactive communication throughout development and review
SESSION 3
Criteria and points of focus: what to have in hand before applying
▶︎
The eligibility criteria for BDD — showing the potential for more effective treatment or diagnosis, and meeting any one of the additional statutory criteria
▶︎
What the reviewing side is actually looking at
▶︎
There is no single prescribed evidence package for a BDD request — what level and type of clinical and non-clinical data strengthens the rationale for designation
SESSION 4
The organization needed to make a BDD submission happen: when to consider it, and who to work with
▶︎
How to choose between a Pre-Submission or another FDA consultation and a BDD
▶︎
How to build the rationale for BDD — clinical need, existing alternatives, supporting evidence, and clinician / KOL input
▶︎
What kind of regulatory advisor to choose, and what experience to ask of them
\🎁 Recording sent to everyone who registers /
Register for the seminar (free)
arrow_upwardWho the seminar is for
✔︎
Executives and business development leads at medical device and IVD manufacturers aiming for approval in the US market
✔︎
Companies planning an FDA consultation (a Pre-Submission or similar) within the next one to three years
✔︎
Regulatory and quality leads who have been through a Pre-Submission and found a single consultation was not enough
✔︎
Those who want to determine whether their own product meets the BDD eligibility criteria — how to argue effectiveness against a life-threatening or irreversibly debilitating condition, plus one of the additional criteria
✔︎
Those considering a BDD submission who want to know what experience to look for in a regulatory advisor
Event details
・Date and time:Tuesday, 27 October 2026, 10:00–11:00 (Japan Standard Time / UTC+9)
・Format:Online (Zoom webinar)
・Fee:Free
・Capacity:200 (planned)
・Registration closes:Monday, 26 October 2026, 23:59 (JST)
・Language:English throughout — slides, presentation and Q&A (no interpretation or subtitles)
・Joining link:emailed to you by xCARE within about 5 minutes of registering, with reminders the day before and one hour before the start
\🎁 Recording sent to everyone who registers /
Register for the seminar (free)
arrow_upwardOn some browsers the registration form may not display correctly. If that happens, please use the link below.
👉 Open the registration form directly